Biotech software development
Custom software for biotech companies, CROs, and research organisations whose laboratory workflows, data volumes, and regulatory obligations require more than generic LIMS platforms or spreadsheet-based tracking can deliver.
Regulatory compliance is designed in from the start: audit trails, electronic signatures, data integrity controls, and the validation documentation FDA and EMA inspectors expect. Not retrofitted after the platform is built.
LIMS with sample lifecycle management, instrument integration, and audit-ready data capture
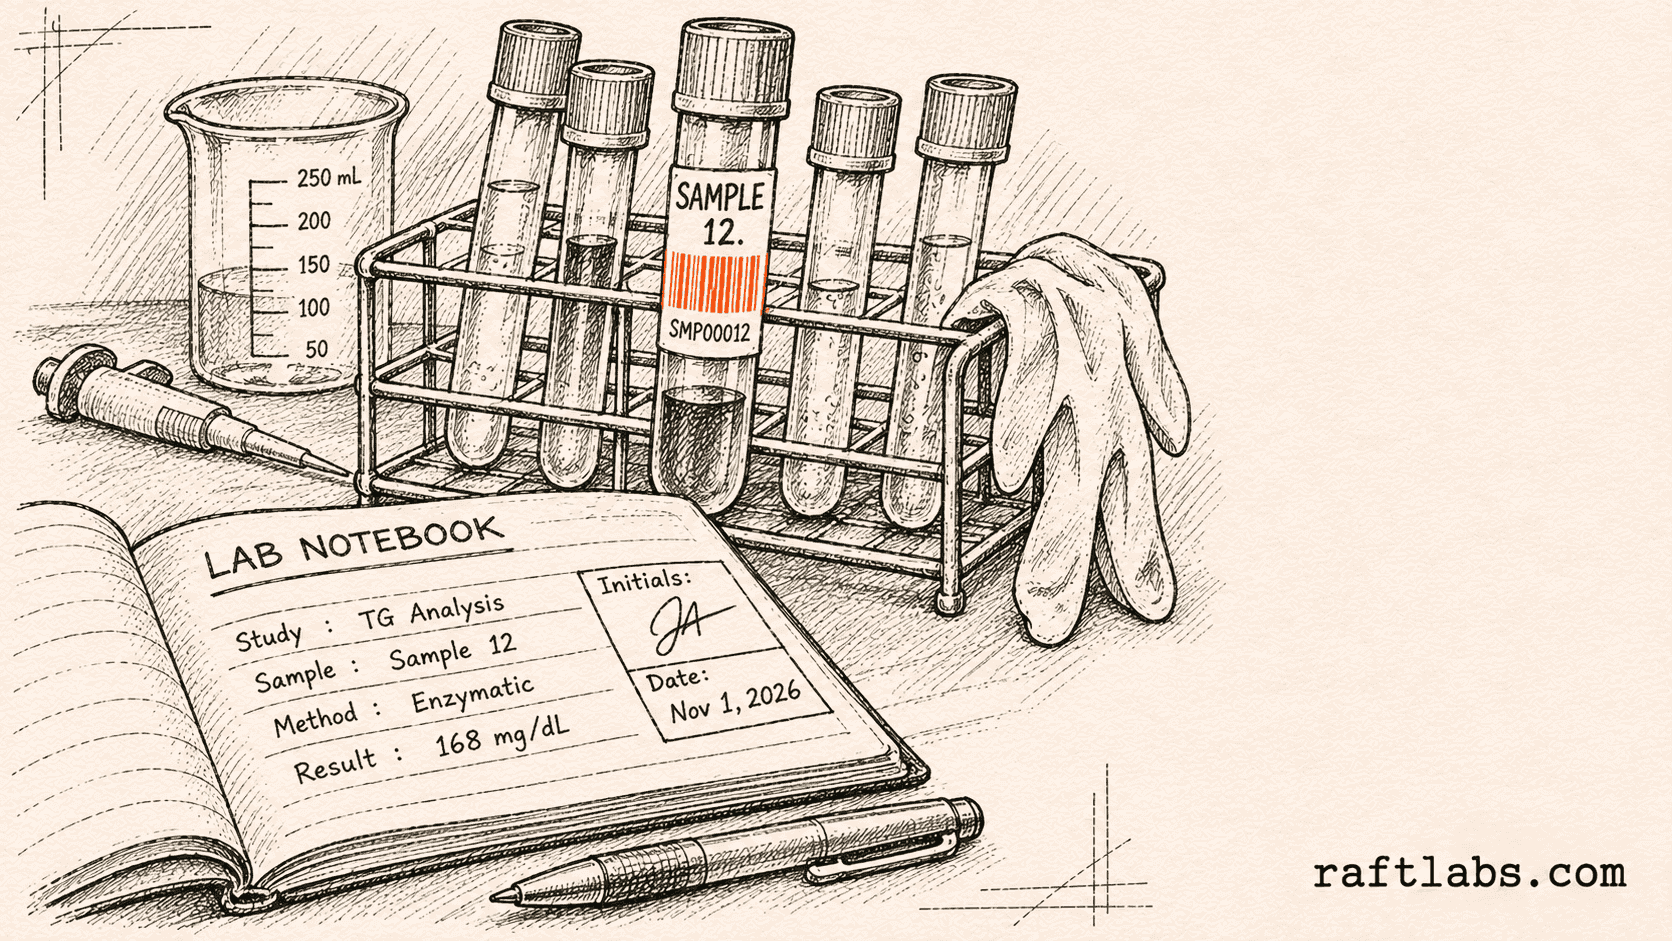
Electronic lab notebooks replacing paper records with structured, searchable, 21 CFR Part 11 compliant data
Clinical trial data management, protocol tracking, subject management, AE reporting, and regulatory submissions
Bioinformatics pipelines for genomics, proteomics, and next-generation sequencing data processing
The problem
Sound familiar?
Research data scattered across instrument exports, spreadsheets, and shared drives with no single source of truth and no audit trail showing who changed what and when?
Regulatory submission preparation consuming weeks of effort to compile the data that should have been structured and traceable from the moment it was generated?
Short answer
RaftLabs builds custom biotech and life sciences software: LIMS, electronic lab notebooks, clinical trial data management, bioinformatics pipelines, and FDA 21 CFR Part 11 compliant systems. Compliance, audit trails, and validation documentation are designed in from the first sprint. Most teams launch a validated v1 in 14 to 20 weeks at a fixed cost of $40K to $110K.
Who this is for
Biotech teams we build for
Biotech and life sciences companies
LIMS, ELN, and data platforms for laboratory workflows that generic systems cannot absorb.
CROs and research organisations
Clinical trial data management, protocol tracking, and submission-ready data capture.
Genomics and bioinformatics teams
Sequencing pipelines and data platforms sized for the storage and compute your work actually needs.
Regulated research labs
21 CFR Part 11 compliant systems with validation documentation scoped from day one.
Regulatory compliance is not a feature you add later in life sciences software
Biotech and life sciences software operates under regulatory frameworks: FDA 21 CFR Part 11, EU Annex 11, and GxP guidelines. All three require audit trails, electronic signatures, access controls, and data integrity protections built into the system architecture from the start. Platforms built without these controls and later validated for regulatory use are expensive to retrofit. They also create ongoing validation burden every time a change is made.
We build life sciences software with the compliance architecture in place from the first sprint. The audit trail captures every data creation, modification, and deletion event with user identity and timestamp. Electronic signatures are built to the 21 CFR Part 11 model: attributable, timestamped, and tamper-evident. Access controls are role-based and documented. The data model is designed against the ALCOA+ principles that FDA and EMA inspectors apply to data integrity: every record attributable, legible, contemporaneous, original, and accurate, and complete, consistent, enduring, and available. The validation approach, IQ, OQ, PQ documentation, is scoped at the start of the project so the system is inspection-ready when it goes live. Where clinical or patient-identifiable data is involved, access controls and audit trails are built with HIPAA-grade protections for that data.
01 Diagnosis
Problems we solve in biotech
- 01Problem
Lab data lives across instrument exports, shared drives, and spreadsheets with no audit trail
CostWhen a scientist needs to work out where a result came from, which instrument produced the raw data, who reviewed it, and what version of the method was in use, that search spans four systems and takes hours. Every hour spent on that search is time not spent on research. For an FDA inspection, that history has to be immediate and complete.
SolutionA LIMS that captures instrument output, reviewer decisions, and method versions in a single record, with an audit trail that cannot be changed, makes that history available in seconds. The investigation that used to take a morning now takes a click.
- 02Problem
Regulatory submission prep takes six to eight weeks because study data was never structured for submission from the start
CostWhen clinical or analytical data is collected in formats that made sense for the study but not for submission, the submission team rebuilds the dataset from scratch. That means free-text fields where coded values (standard codes, not free text) are required, patient identifiers where anonymisation is needed, and result formats that don't map to CDISC SDTM. Every week of that effort is a week between study completion and IND or NDA filing.
SolutionA data capture system designed around submission requirements from the start produces submission-ready output as a side effect of normal operations. Filing prep drops from weeks to days.
- 03Problem
Lab notebooks are physical binders that can't be searched, shared between sites, or countersigned electronically for IP protection
CostWhen research is recorded on paper, a scientist at a second site can't access the notebook. A colleague can't countersign it without being in the same room. A patent attorney can't search it without physically reviewing every page. If a priority date dispute arises (a dispute over who did the work first), the documentation is slow to produce and hard to verify.
SolutionAn electronic lab notebook with structured entries, electronic countersignature, and full-text search makes the research record immediately accessible and legally defensible.
- 04Problem
Your LIMS tracks samples but has no instrument integration, so technicians re-enter results by hand from printouts
CostWhen instrument results are printed, checked by a technician, and manually entered into the LIMS, that transcription step (copying the result by hand) is the most common source of data integrity failure in laboratory operations. A 2024 systematic review of data-entry accuracy in clinical research (indexed in PubMed) found that single-entry manual data processing carries a pooled error rate of 0.29%, with high variability, reaching as many as 2,784 errors per 10K data fields in the worst-performing manual methods. An instrument that produces 200 results per day creates 200 opportunities for a digit to be entered incorrectly, and one wrong value can invalidate a batch or trigger a regulatory query.
SolutionDirect instrument integration, via API, ASTM interface, or file-based import, removes the transcription step entirely. It also creates a direct link between the raw instrument output and the LIMS result record so the chain of custody is unbroken.
02 What we ship
What we build
Laboratory information management system (LIMS)
Sample registration and tracking through every stage of the laboratory workflow: receipt, storage, testing, review, and disposal, with sample status and location visible at all times. Test order management links samples to analytical methods and instrument assignments. Results are captured via direct instrument integrations and manual entry, with units, ranges, and pass/fail criteria per test type. Review and approval runs through an electronic signature workflow. Certificates of analysis generate automatically from approved results. Your laboratory gets the data traceability a regulatory inspection requires, without the paper worksheets.
Electronic lab notebook (ELN)
Structured experiment records replace paper lab notebooks with searchable, version-controlled, electronically signed entries. Experiment templates capture the protocol, materials, equipment, and observations in a consistent format every time. Instrument exports, images, chromatograms, and spectra attach directly to the experiment record. Multi-researcher experiments track contributions individually. Timestamped, countersigned entries establish priority and demonstrate inventive step for IP protection. Your research data becomes findable, shareable, and defensible without leaving the system.
Clinical trial data management
Protocol management captures study design, endpoints, visit schedule, and inclusion/exclusion criteria. Subject enrolment and randomisation run eligibility checks against the protocol automatically. Electronic data capture handles case report form completion with field-level validation and query management. Adverse event and SAE reporting follow a regulatory notification workflow. Site management covers investigator files, monitoring visit records, and deviation tracking. Clinical data exports in CDISC SDTM and ADaM format for regulatory submission. Your study stays in protocol and your submission data arrives structured for FDA review.
Bioinformatics pipeline platform
Next-generation sequencing data processing covers alignment, variant calling, annotation, and quality control, with configurable workflow steps and parameter management. Genomics data storage includes sample linkage, metadata management, and version control for reference data. Pipeline execution management shows run status, resource usage, and output file tracking in one view. Results visualisation covers variant data, expression profiles, and pathway analysis. Controlled access data sharing supports collaboration between internal and external research teams. Raw sequencing output becomes analysed, reproducible, shareable results.
FDA 21 CFR Part 11 compliance
Audit trail captures every record creation, modification, and deletion with user identity, timestamp, and previous value: the complete change history that 21 CFR Part 11 and EU Annex 11 require. Electronic signature workflow includes identity verification, meaning capture, and binding of the signature to the record. Access control covers role-based permissions, password policy enforcement, and inactivity timeout. System validation documentation, User Requirements Specification, Functional Specification, IQ/OQ/PQ protocols, and Validation Summary Report, is produced alongside the software. Change control procedure for post-validation modifications maintains the validated state throughout.
Regulatory data and submission tools
Data compilation tools aggregate study data, analytical results, and supporting documentation into the structured format required for IND, NDA, BLA, or MAA submissions. eCTD document management covers module structure, document versioning, and submission package assembly. Regulatory correspondence tracking includes response timelines and commitment management. Post-approval change control documentation runs through a defined workflow. Submission data becomes a structured output of normal operations, not a manual compilation exercise. The time between study completion and submission-ready drops measurably.
Integrations
Works with the systems you already run
Instrument integration and submission formats are scoped in discovery against your specific equipment and requirements.
- Instrument interface standards
- ASTM · HL7 · CSV and XML instrument exports
- Submission standards
- CDISC SDTM · CDISC ADaM · eCTD
- Cloud infrastructure
- AWS · Azure · GCP
03 Buy, build, or wait
Stay on a configured platform while your workflows are standard and stable
The biotech software question is really about whether your lab workflows and regulatory obligations fit inside a generic LIMS.
Buy or rent
A configured LIMS, for standard lab workflows
When sample tracking, test management, and reporting fit the vendor's data model without workarounds.
Wait, while the lab process is still changing
When protocols and instrument setups are in flux, lock the workflow before building around it.
Build custom
When 21 CFR Part 11 must be designed in
Audit trails, electronic signatures, and validation documentation from the first sprint.
When instruments need direct integration
API, ASTM, or file-based capture that removes manual transcription entirely.
When submission-ready data is the goal
Data captured in CDISC-ready structure as a side effect of daily operations.
Bottom line
Stay on a configured platform while your workflows are standard and stable. Build when compliance, instruments, or data structure demand it.
04 How we work
How we work with biotech clients
- Step 0101
Discovery
We map your laboratory workflows, regulatory obligations (21 CFR Part 11, EU Annex 11, GxP guidelines), and the instrument types requiring integration before any design begins. For clinical trial projects, we document the study protocol structure, data capture requirements, and CDISC submission format requirements at this stage.
- Step 0202
Architecture
We design the data model with regulatory compliance as a first-class constraint: audit trail schema, electronic signature binding, role-based access controls, and the validation documentation plan. Instrument integration approaches are confirmed based on available APIs and interface protocols for your specific equipment.
- Step 0303
Build
Two-week sprints with working software at each checkpoint. Core sample tracking or data capture functionality ships first. Instrument integrations, review workflows, electronic signatures, and reporting follow in subsequent sprints, with IQ/OQ protocol documentation written in parallel.
- Step 0404
Launch and support
Performance Qualification runs with your validation team against your specific processes. The Validation Summary Report and supporting documentation are delivered alongside the live system. Post-launch support covers regulatory updates, new instrument integrations, and change control documentation for post-validation modifications.
05 Track record
What biotech teams get when they work with us
- Validation documentation from sprint one
- Audit-ready
- Validation documentation delivered with the software
- IQ/OQ/PQ
- Cost agreed before development starts
- Fixed
- To a validated v1, then iterate
- 14-20 wks
08 Why us
Why choose us?
- 01
Compliance is designed in, not retrofitted
Audit trails, electronic signatures, and validation documentation are scoped at kickoff and built from the first sprint, so your system is inspection-ready when it goes live. - 02
Instrument integration is confirmed before it is priced
We assess the integration approach for each instrument during discovery and confirm what is achievable for your specific equipment before development starts. No mid-build surprises. - 03
Fixed cost, validation documentation included
The URS, Functional Specification, IQ/OQ protocols, and Validation Summary Report are delivered alongside the software. Cost is fixed and agreed before work starts.
Frequently asked questions
Straight answers to what buyers ask before they scope. Still stuck? .
Validation is scoped as part of the development project, not treated as a separate post-development activity. We produce the User Requirements Specification and Functional Specification during the discovery phase so requirements are documented before development starts. Installation Qualification and Operational Qualification protocols are written in parallel with development and executed during the testing phase. Performance Qualification runs with your validation team against your specific processes. The Validation Summary Report and supporting documentation are delivered alongside the software. The validation approach is discussed and agreed at project kickoff so the timeline and deliverables are clear from day one.
Yes. Instrument integration is a core part of LIMS implementation. We support direct API integration where instruments provide it, bi-directional instrument interfaces via ASTM or HL7 protocols, file-based import from instrument software exports (CSV, XML, proprietary formats), and middleware integration platforms. The integration approach for each instrument is assessed during discovery. Not all instruments offer equivalent integration capability, and we confirm what's achievable for your specific equipment before development starts.
Genomics data is large by nature and the storage and compute architecture needs to be designed for it from the start. We build bioinformatics platforms on cloud infrastructure (AWS, Azure, or GCP) with elastic storage and on-demand compute for pipeline execution, so you pay for the capacity you use rather than provisioning for peak. Data archiving workflows manage the lifecycle of raw sequencing files, intermediate files, and final outputs. We assess your anticipated data volume and pipeline complexity during discovery and propose the right infrastructure architecture before development starts.
A validated v1 costs $40K to $110K: an LIMS, electronic lab notebook, or bioinformatics module takes 14 to 20 weeks. A full multi-module platform costs $120K to $400K. Validation documentation is included in the project scope and cost. We scope every project before pricing, and the cost is fixed and agreed before work starts.
Yes. Data migration of historical laboratory records, paper worksheets, Excel exports, legacy LIMS exports, is a standard part of biotech software projects. We structure the migration around sample identifiers, analytical methods, and result records so the new system starts with an accurate historical baseline. For paper records, we scope a structured data capture process as part of migration. The migration approach is confirmed during discovery so the timeline and quality controls are clear before development starts.


Biotech software by product
- 01
AI document intelligence
Extract structured data from clinical trial reports, lab results, regulatory submissions, and audit trails
- 02
AI agent development
Autonomous agents for literature review, protocol deviation monitoring, and regulatory document generation
- 03
Business process automation
Automate sample tracking workflows, equipment calibration scheduling, and regulatory reporting
- 04
Custom software development
Custom LIMS, ELN, and clinical data management platforms built to your GMP/21 CFR Part 11 requirements
Get a build-vs-buy plan for your team.
Tell us your laboratory workflows, your regulatory obligations, and where your current systems create compliance risk or research inefficiency. We'll scope the right platform and give you a fixed cost.
- Scope and cost agreed before work starts. No surprises. No obligation.
- Working prototype within 3 weeks of kickoff.
- Pay by milestone. You see progress before each invoice.
- 60-day post-launch warranty. Bug fixes, UI tweaks, and deployment support. No retainer.
- All conversations are NDA-protected.